









| 查看: 9362 | 回复: 2 | |||
| 【悬赏金币】回答本帖问题,作者lm20220613将赠送您 5 个金币 | |||
[求助]
二氧化铈功函数计算求助 已有1人参与
|
|||

请教大家:我用VASP计算二氧化铈(200)晶面的功函数,表面静电势结果如下,波动峰的高度不一致,这是为啥呢?还有,我加了偶极矫正,但是真空能级的位置还是向下倾斜。我附上了我的INCAR和晶胞结构,烦请大家帮我看看,非常感谢大家 。 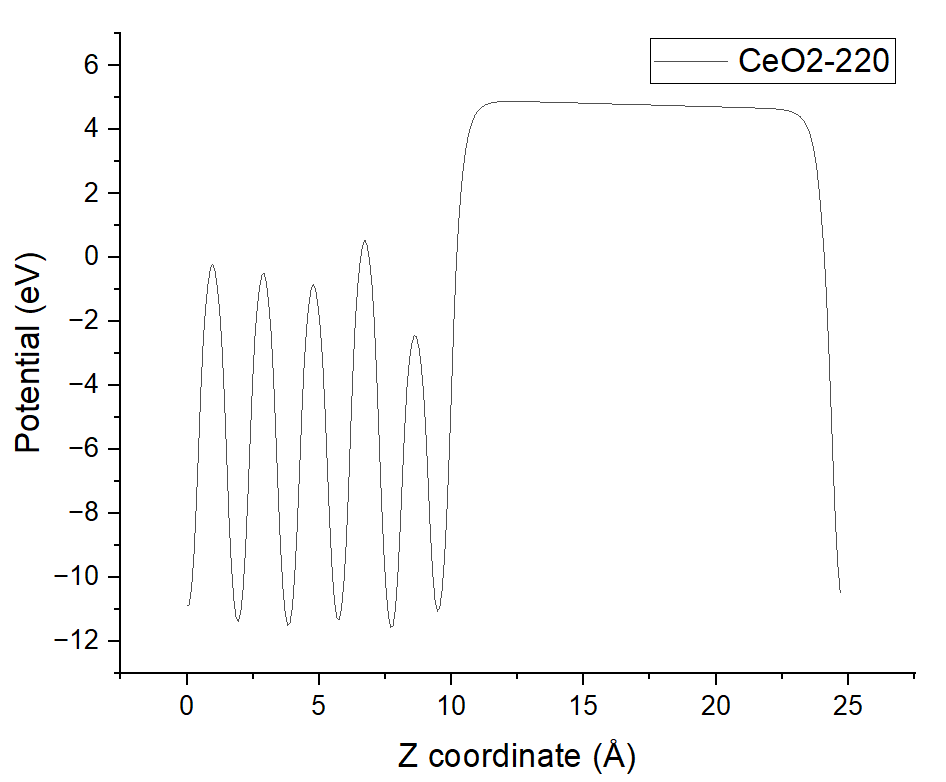 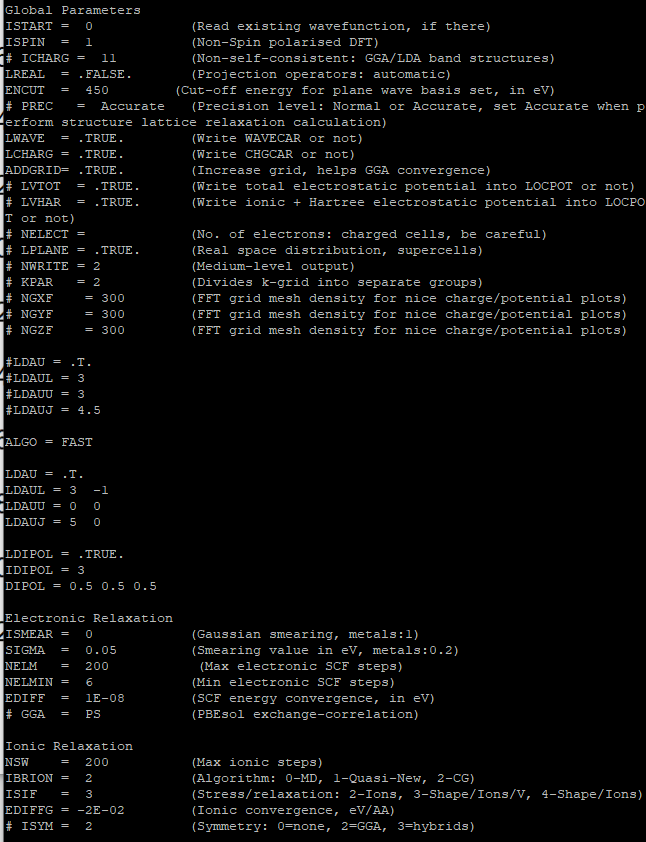 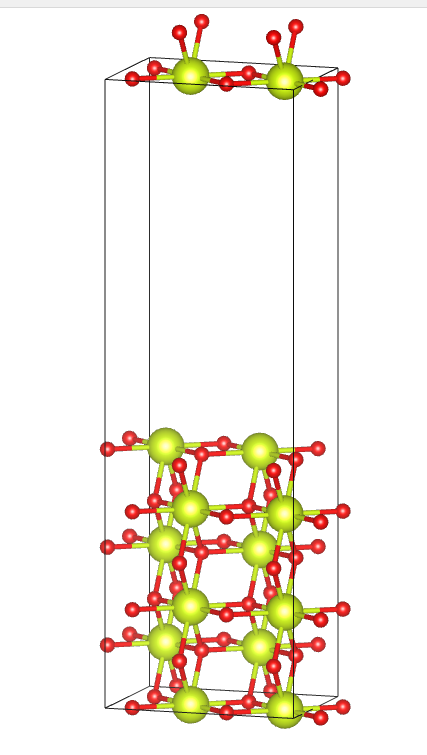 |
 回复此楼
» 猜你喜欢
 2027博士申请-临床检验诊断学、病原生物学、微生物方向
已经有5人回复
-
【长期招聘】复旦大学 AI 制药实验室 博士后招聘
已经有10人回复
-
各位基础老师们,看完中央《从严治党》的决定后,对学术界更加有信心了吧
已经有4人回复
-
越来越担心硕士毕业后没有竞争力了
已经有15人回复
-
申博自荐 2篇一区TOP+1篇EI+2公开发明专利 2027博士申请
已经有3人回复
HER12025
铁虫 (小有名气)
- 应助: 0 (幼儿园)
- 金币: 231.3
- 散金: 35
- 帖子: 213
- 在线: 120.3小时
- 虫号: 36722173
- 注册: 2025-12-26
- 专业: 高分子材料结构与性能

2楼2026-01-06 16:38:11
【答案】应助回帖
|
Your slab model contains roughly 5 atomic planes. For cerium dioxide ($\text{CeO}_2$), this is quite thin. Without a thick enough "bulk core," the structural relaxations from both surfaces propagate through the entire slab. The interlayer distances and planar charge densities vary from layer to layer, preventing the electrostatic potential from stabilizing into a perfectly periodic oscillation. By default, VASP's dipole correction algorithm expects the slab to be continuous in the center of the cell and the vacuum to be at the boundaries ($Z=0$ and $Z=c$). Alternatively, if it evaluates the center of charge automatically without an explicit DIPOL tag, the split nature of your slab misleads the algorithm. The potential step/jump intended to cancel the dipole field is likely being placed in the wrong position potentially overlapping with the atoms at the top of your box ($Z > 24\text{ \AA}$), which causes the sharp drop-off and leaves an uncompensated field in the vacuum. Modify your POSCAR so that the entire slab is continuous and centered in the middle of the simulation box (around $Z = c/2$). The vacuum should be split evenly at the bottom ($Z = 0$) and the top ($Z = c$). This allows VASP to easily identify the vacuum region and properly apply the dipole step function. |
3楼2026-05-20 17:26:26